
MXenes are a rapidly growing family of ultra-thin, two-dimensional materials that show promise for applications ranging from batteries and sensors to electronics, catalysts, and renewable energy technologies. One of the biggest challenges facing researchers is that MXenes can be made from many different combinations of metals and surface chemistries, making it difficult to predict which compositions will deliver the best performance for a particular application. Most previous studies examined only a small number of MXenes, leaving much of the material design space unexplored.
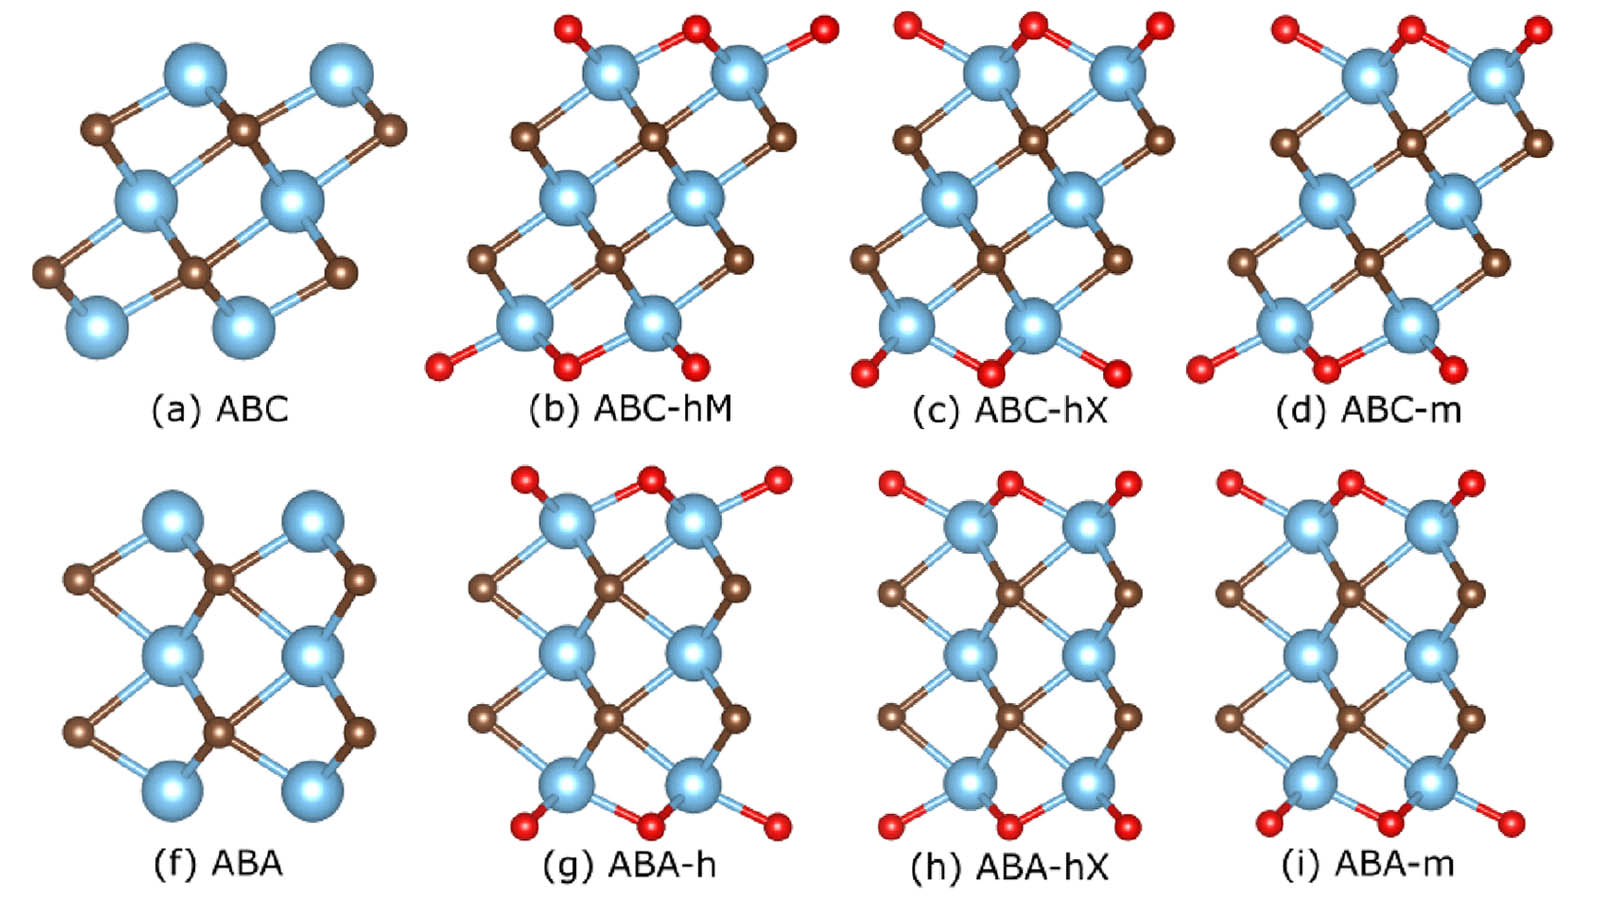
In this study, researchers Emily Sutherland, Benjamin Traverso, and N. Aaron Deskins conducted one of the most comprehensive computational investigations of MXenes to date. Using more than 2,000 quantum-mechanical simulations, they examined 99 different M₃C₂T₂ MXenes containing a wide range of metals and surface terminations. The team discovered that composition strongly influences a material's structure, electrical behavior, and mechanical properties, and they identified numerous previously unexplored MXenes with particularly promising characteristics. The study also revealed that about 20% of the materials preferred a different atomic stacking arrangement than commonly assumed in earlier modeling studies.
The findings provide a valuable roadmap for scientists developing next-generation materials for energy storage, catalysis, sensing, and electronic devices. Rather than relying on trial and error in the laboratory, researchers can use these results to focus on the most promising candidates. Research computing was central to the project, enabling large-scale density functional theory (DFT) calculations that would be impractical through experimentation alone and allowing the team to systematically map how composition determines MXene properties.
Sutherland, Emily et al (2025), Impact of composition on the structural, electronic, and mechanical properties of M3C2T2 MXenes, Mater. Adv., doi: 10.1039/d5ma00874c